Company CEO outlines the differentiated mechanism and clinical potential of Ampligen in pancreatic cancer

Survival data highlighted in accompanying chart show Ampligen-associated median overall survival of 34.8 months in a selected pancreatic patient population versus 12.5 months in historical controls

OCALA, Fla., Sept. 22, 2026 (GLOBE NEWSWIRE) — AIM ImmunoTech Inc. (NYSE American: AIM) (“AIM” or the “Company”) today issued a letter to stockholders from Chief Executive Officer Thomas Equels addressing the recent U.S. Food and Drug Administration (the “FDA”) approval of daraxonrasib and outlining the Company’s view of Ampligen’s differentiated mechanism, clinical profile and potential role in the future treatment of pancreatic cancer.

The full letter follows:

Dear Fellow Stockholders,

The FDA’s recent approval of daraxonrasib is meaningful progress for people with metastatic pancreatic cancer. In fact, it is significant enough that you may be asking – and I should answer – two questions:

Question 1: How is AIM’s drug Ampligen® different from daraxonrasib?

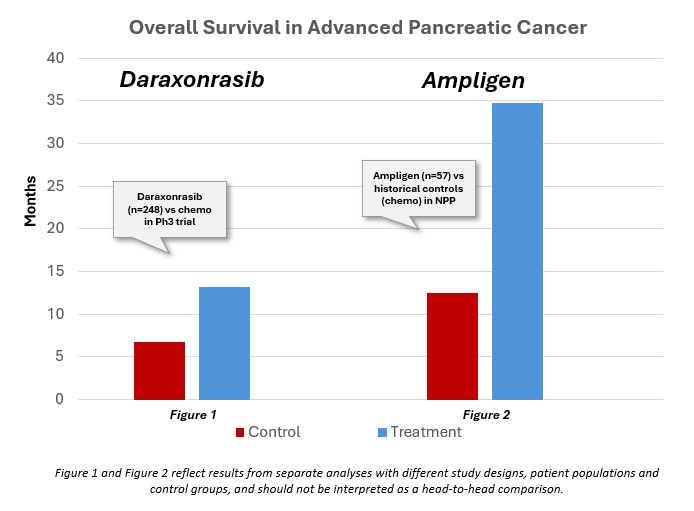
Daraxonrasib is a selective, targeted therapy that inhibits RAS, a major driver of pancreatic cancer. A pivotal, randomized, well-controlled study demonstrated median overall survival of 13.2 months with daraxonrasib compared to 6.7 months with standard chemotherapy, for an improvement of 6.5 months (See Figure 1 below, NEJM). These results are an important part of the basis for the FDA’s approval of daraxonrasib, but they also indicate that many patients on both daraxonrasib and on standard therapy experience disease progression and mortality. Further, in laboratory studies, pancreatic tumors have been seen to develop resistance to RAS inhibition.

Ampligen has a different mechanism of action from daraxonrasib. We believe that Ampligen is the only clinically advanced investigational TLR3 agonist with a well-developed safety profile that is designed to activate innate immunity and help therapeutically reshape the tumor microenvironment to provide a natural mechanism to eliminate cancer cells. While daraxonrasib directly inhibits one driver mutation related to pancreatic cancer, Ampligen has the potential to induce a broad-spectrum therapeutic effect that amplifies a patient’s innate immune system, with data suggesting the potential for consequent stabilization of the pancreatic cancer tumor immune responses with improvement in a patient’s Quality of Life. We have seen evidence of this therapeutic potential not only in pancreatic cancer, but in a range of other solid tumors.

Question 2: What role will Ampligen serve in the future of pancreatic cancer care?

Daraxonrasib is an important therapeutic advance, but pancreatic cancer is not a simple disease where one successful targeted drug makes additional therapeutic development unnecessary. We believe Ampligen has the potential to complement and extend therapies such as daraxonrasib by helping to overcome the immune biological barriers that have historically limited treatment success in pancreatic cancer.

Further, we believe that Ampligen has significant potential in pancreatic cancer as a monotherapy. In an analysis of patients with a blood neutrophil-to-lymphocyte ratio below 4.5 who participated in a Dutch government-approved Named Patient Program, Ampligen as a monotherapy was linked to a median overall survival of 34.8 months compared with only 12.5 months in well-matched historical controls, representing an observed increase in median overall survival of 22.3 months over the historical controls (See Figure 2 below, AIMImmuno.com).
Screenshot 2026-09-21

The accompanying chart visually summarizes the respective overall-survival results cited in this letter, including the separate control and treatment comparisons for daraxonrasib and Ampligen based on the separate cited datasets.

In conclusion, daraxonrasib’s approval does not preclude a potentially important role for Ampligen. Daraxonrasib targets a single driver of pancreatic cancer, while Ampligen is designed to work by activating and enhancing the body’s innate immune system to target multiple types of solid tumors, including cancers of the pancreas. Based on all that we know to date, we believe that Ampligen continues to have an important potential role to serve in pancreatic cancer treatment.

AIM is also invested in Ampligen’s potential as a part of a combination therapy with checkpoint inhibitors in a broad range of other solid tumors, including pancreatic cancer. The ongoing Phase 2 DURIPANC clinical trial is evaluating Ampligen with AstraZeneca’s durvalumab (Imfinzi) in patients whose pancreatic cancer was stable post-FOLFIRINOX. This is an exploratory, open-label, single-center study with a small number of patients. Its primary endpoint is the proportion of patients who have stable disease or a tumor response 24 weeks after combination treatment begins. We anticipate topline results from the DURIPANC study in Q1 2027, followed by a detailed analysis of overall survival in Q3 2027. AIM believes that Ampligen’s observed immune activity will translate into a meaningful clinical signal and allow us to establish the parameters of a follow-up, pivotal Phase 3 clinical trial.

While there is still work to be done, including rigorous pivotal trials, our goal remains clear: to determine whether Ampligen can contribute to longer and better lives for patients with pancreatic cancer. We believe it can.

AIM appreciates your continued support as we pursue that goal with scientific discipline and a clear understanding of what still must be proven.

Thomas K. Equels
Chief Executive Officer
AIM ImmunoTech Inc.

References:

About AIM ImmunoTech Inc.

AIM ImmunoTech Inc. is an immuno-pharma company focused on the research and development of its lead product, Ampligen® (rintatolimod), for the treatment of late-stage pancreatic cancer, a lethal and unmet global health problem. Ampligen is a dsRNA and highly selective TLR3 agonist immuno-modulator that has shown broad-spectrum activity in clinical trials.

For more information, please visit aimimmuno.com and connect with the Company on X, LinkedIn, and Facebook.

Cautionary Statement

This press release contains forward-looking statements within the meaning of Section 27A of the Securities Act of 1933, as amended, and Section 21E of the Securities Exchange Act of 1934, as amended, that involve a number of risks and uncertainties. Any forward-looking statements set forth in this press release speak only as of the date hereof. Such forward-looking statements may include statements relating to: Ampligen’s potential to reshape the pancreatic tumor microenvironment and to enhance the activity of PD-1 and PD-L1 checkpoint inhibitors, including pembrolizumab and durvalumab; the potential for any such effect to be durable or long-lasting; Ampligen’s potential to complement existing therapies and to overcome biological barriers that have limited treatment success in pancreatic cancer; the potential of the Phase 2 DURIPANC study to inform patient selection, biomarker identification and the design of a potential future pivotal development program; the timing, conduct and outcome of the DURIPANC primary endpoint analyses, biomarker evaluations and overall survival follow-up; the timing and outcome of planned regulatory interactions; intellectual property expansion; the Company’s long-term pancreatic cancer development strategy and its efforts to position Ampligen for the next stage of development; and the Company’s ability to create long-term value for its stockholders. Forward-looking statements include statements attributed to, or made by, the Company’s Chief Executive Officer in the letter above. For all forward-looking statements, the Company claims the protection of the safe harbor for forward-looking statements contained in the Private Securities Litigation Reform Act of 1995. All statements in this press release other than statements of historical fact, including statements regarding the potential of Ampligen’s mechanism of action, anticipated regulatory milestones, our future results of operations and financial position, our business strategy and plans, and our objectives for future operations, are forward-looking statements. Words such as “believe,” “may,” “might,” “will,” “could,” “should,” “can,” “estimate,” “continue,” “anticipate,” “intend,” “expect,” “envision,” “potential” and similar expressions are intended to identify forward-looking statements, but the absence of these words does not mean that a statement is not forward-looking. The Company does not undertake to update any of these forward-looking statements to reflect events or circumstances that occur after the date hereof, except as required by applicable law. The Company is in various stages of seeking to determine whether Ampligen will be effective in the treatment of multiple types of viral diseases, cancers, and immune-deficiency disorders, and disclosures in the Company’s reports filed with the SEC, on its website, and in its press releases set forth its current and anticipated future activities. These activities are subject to change for a number of reasons. Significant additional testing and trials will be required to determine whether Ampligen® will be effective in the treatment of these conditions. Results obtained in preclinical studies do not necessarily predict results in humans. Human clinical trials will be necessary to prove whether or not Ampligen® will be efficacious in humans. No assurance can be given as to whether current or planned clinical trials will be successful or yield favorable data, and the trials are subject to many factors including lack of regulatory approval(s), lack of study drug, lack of adequate funding, or a change in priorities at the institutions sponsoring other trials. Even if these clinical trials are initiated, the Company cannot assure that the clinical studies will be successful or yield any useful data. No assurance can be given that the findings in preliminary studies will prove true or that such studies will yield favorable results, or that future studies will not result in findings that are different from those reported in the studies referenced in the Company’s reports filed with the SEC, on the Company’s website, and in its press releases. Operating in foreign countries carries with it a number of risks, including potential difficulties in enforcing intellectual property rights. The Company cannot assure that its potential foreign operations will not be adversely affected by these risks.

For a detailed discussion of these and other risk factors, please review the “Risk Factors” section in the Company’s most recent Annual Report on Form 10-K and subsequent Quarterly Reports on Form 10-Q filed with the U.S. Securities and Exchange Commission. These filings are available at www.sec.gov and www.aimimmuno.com. You should not place undue reliance on any forward-looking statements. The information found on the Company’s website or on other websites referenced or linked to in this press release is not incorporated by reference into this press release and such information is referenced or linked for reference purposes only.

A photo accompanying this announcement is available at https://www.globenewswire.com/NewsRoom/AttachmentNg/53bfb7cd-009a-4cc0-9b40-c044efef4b9c

CONTACT: Investor Contact:

JTC Team, LLC
Jenene Thomas
908.824.0775
AIM@jtcir.com

Company CEO outlines the differentiated mechanism and clinical potential of Ampligen in pancreatic cancer

Survival data highlighted in accompanying chart show Ampligen-associated median overall survival of 34.8 months in a selected pancreatic patient population versus 12.5 months in historical controls

OCALA, Fla., Sept. 22, 2026 (GLOBE NEWSWIRE) — AIM ImmunoTech Inc. (NYSE American: AIM) (“AIM” or the “Company”) today issued a letter to stockholders from Chief Executive Officer Thomas Equels addressing the recent U.S. Food and Drug Administration (the “FDA”) approval of daraxonrasib and outlining the Company’s view of Ampligen’s differentiated mechanism, clinical profile and potential role in the future treatment of pancreatic cancer.

The full letter follows:

Dear Fellow Stockholders,

The FDA’s recent approval of daraxonrasib is meaningful progress for people with metastatic pancreatic cancer. In fact, it is significant enough that you may be asking – and I should answer – two questions:

Question 1: How is AIM’s drug Ampligen® different from daraxonrasib?

Daraxonrasib is a selective, targeted therapy that inhibits RAS, a major driver of pancreatic cancer. A pivotal, randomized, well-controlled study demonstrated median overall survival of 13.2 months with daraxonrasib compared to 6.7 months with standard chemotherapy, for an improvement of 6.5 months (See Figure 1 below, NEJM). These results are an important part of the basis for the FDA’s approval of daraxonrasib, but they also indicate that many patients on both daraxonrasib and on standard therapy experience disease progression and mortality. Further, in laboratory studies, pancreatic tumors have been seen to develop resistance to RAS inhibition.

Ampligen has a different mechanism of action from daraxonrasib. We believe that Ampligen is the only clinically advanced investigational TLR3 agonist with a well-developed safety profile that is designed to activate innate immunity and help therapeutically reshape the tumor microenvironment to provide a natural mechanism to eliminate cancer cells. While daraxonrasib directly inhibits one driver mutation related to pancreatic cancer, Ampligen has the potential to induce a broad-spectrum therapeutic effect that amplifies a patient’s innate immune system, with data suggesting the potential for consequent stabilization of the pancreatic cancer tumor immune responses with improvement in a patient’s Quality of Life. We have seen evidence of this therapeutic potential not only in pancreatic cancer, but in a range of other solid tumors.

Question 2: What role will Ampligen serve in the future of pancreatic cancer care?

Daraxonrasib is an important therapeutic advance, but pancreatic cancer is not a simple disease where one successful targeted drug makes additional therapeutic development unnecessary. We believe Ampligen has the potential to complement and extend therapies such as daraxonrasib by helping to overcome the immune biological barriers that have historically limited treatment success in pancreatic cancer.

Further, we believe that Ampligen has significant potential in pancreatic cancer as a monotherapy. In an analysis of patients with a blood neutrophil-to-lymphocyte ratio below 4.5 who participated in a Dutch government-approved Named Patient Program, Ampligen as a monotherapy was linked to a median overall survival of 34.8 months compared with only 12.5 months in well-matched historical controls, representing an observed increase in median overall survival of 22.3 months over the historical controls (See Figure 2 below, AIMImmuno.com).

Screenshot 2026-09-21

The accompanying chart visually summarizes the respective overall-survival results cited in this letter, including the separate control and treatment comparisons for daraxonrasib and Ampligen based on the separate cited datasets.

In conclusion, daraxonrasib’s approval does not preclude a potentially important role for Ampligen. Daraxonrasib targets a single driver of pancreatic cancer, while Ampligen is designed to work by activating and enhancing the body’s innate immune system to target multiple types of solid tumors, including cancers of the pancreas. Based on all that we know to date, we believe that Ampligen continues to have an important potential role to serve in pancreatic cancer treatment.

AIM is also invested in Ampligen’s potential as a part of a combination therapy with checkpoint inhibitors in a broad range of other solid tumors, including pancreatic cancer. The ongoing Phase 2 DURIPANC clinical trial is evaluating Ampligen with AstraZeneca’s durvalumab (Imfinzi) in patients whose pancreatic cancer was stable post-FOLFIRINOX. This is an exploratory, open-label, single-center study with a small number of patients. Its primary endpoint is the proportion of patients who have stable disease or a tumor response 24 weeks after combination treatment begins. We anticipate topline results from the DURIPANC study in Q1 2027, followed by a detailed analysis of overall survival in Q3 2027. AIM believes that Ampligen’s observed immune activity will translate into a meaningful clinical signal and allow us to establish the parameters of a follow-up, pivotal Phase 3 clinical trial.

While there is still work to be done, including rigorous pivotal trials, our goal remains clear: to determine whether Ampligen can contribute to longer and better lives for patients with pancreatic cancer. We believe it can.

AIM appreciates your continued support as we pursue that goal with scientific discipline and a clear understanding of what still must be proven.

Thomas K. Equels
Chief Executive Officer
AIM ImmunoTech Inc.

References:

About AIM ImmunoTech Inc.

AIM ImmunoTech Inc. is an immuno-pharma company focused on the research and development of its lead product, Ampligen® (rintatolimod), for the treatment of late-stage pancreatic cancer, a lethal and unmet global health problem. Ampligen is a dsRNA and highly selective TLR3 agonist immuno-modulator that has shown broad-spectrum activity in clinical trials.

For more information, please visit aimimmuno.com and connect with the Company on X, LinkedIn, and Facebook.

Cautionary Statement

This press release contains forward-looking statements within the meaning of Section 27A of the Securities Act of 1933, as amended, and Section 21E of the Securities Exchange Act of 1934, as amended, that involve a number of risks and uncertainties. Any forward-looking statements set forth in this press release speak only as of the date hereof. Such forward-looking statements may include statements relating to: Ampligen’s potential to reshape the pancreatic tumor microenvironment and to enhance the activity of PD-1 and PD-L1 checkpoint inhibitors, including pembrolizumab and durvalumab; the potential for any such effect to be durable or long-lasting; Ampligen’s potential to complement existing therapies and to overcome biological barriers that have limited treatment success in pancreatic cancer; the potential of the Phase 2 DURIPANC study to inform patient selection, biomarker identification and the design of a potential future pivotal development program; the timing, conduct and outcome of the DURIPANC primary endpoint analyses, biomarker evaluations and overall survival follow-up; the timing and outcome of planned regulatory interactions; intellectual property expansion; the Company’s long-term pancreatic cancer development strategy and its efforts to position Ampligen for the next stage of development; and the Company’s ability to create long-term value for its stockholders. Forward-looking statements include statements attributed to, or made by, the Company’s Chief Executive Officer in the letter above. For all forward-looking statements, the Company claims the protection of the safe harbor for forward-looking statements contained in the Private Securities Litigation Reform Act of 1995. All statements in this press release other than statements of historical fact, including statements regarding the potential of Ampligen’s mechanism of action, anticipated regulatory milestones, our future results of operations and financial position, our business strategy and plans, and our objectives for future operations, are forward-looking statements. Words such as “believe,” “may,” “might,” “will,” “could,” “should,” “can,” “estimate,” “continue,” “anticipate,” “intend,” “expect,” “envision,” “potential” and similar expressions are intended to identify forward-looking statements, but the absence of these words does not mean that a statement is not forward-looking. The Company does not undertake to update any of these forward-looking statements to reflect events or circumstances that occur after the date hereof, except as required by applicable law. The Company is in various stages of seeking to determine whether Ampligen will be effective in the treatment of multiple types of viral diseases, cancers, and immune-deficiency disorders, and disclosures in the Company’s reports filed with the SEC, on its website, and in its press releases set forth its current and anticipated future activities. These activities are subject to change for a number of reasons. Significant additional testing and trials will be required to determine whether Ampligen® will be effective in the treatment of these conditions. Results obtained in preclinical studies do not necessarily predict results in humans. Human clinical trials will be necessary to prove whether or not Ampligen® will be efficacious in humans. No assurance can be given as to whether current or planned clinical trials will be successful or yield favorable data, and the trials are subject to many factors including lack of regulatory approval(s), lack of study drug, lack of adequate funding, or a change in priorities at the institutions sponsoring other trials. Even if these clinical trials are initiated, the Company cannot assure that the clinical studies will be successful or yield any useful data. No assurance can be given that the findings in preliminary studies will prove true or that such studies will yield favorable results, or that future studies will not result in findings that are different from those reported in the studies referenced in the Company’s reports filed with the SEC, on the Company’s website, and in its press releases. Operating in foreign countries carries with it a number of risks, including potential difficulties in enforcing intellectual property rights. The Company cannot assure that its potential foreign operations will not be adversely affected by these risks.

For a detailed discussion of these and other risk factors, please review the “Risk Factors” section in the Company’s most recent Annual Report on Form 10-K and subsequent Quarterly Reports on Form 10-Q filed with the U.S. Securities and Exchange Commission. These filings are available at www.sec.gov and www.aimimmuno.com. You should not place undue reliance on any forward-looking statements. The information found on the Company’s website or on other websites referenced or linked to in this press release is not incorporated by reference into this press release and such information is referenced or linked for reference purposes only.

A photo accompanying this announcement is available at https://www.globenewswire.com/NewsRoom/AttachmentNg/53bfb7cd-009a-4cc0-9b40-c044efef4b9c

CONTACT: Investor Contact:

JTC Team, LLC
Jenene Thomas
908.824.0775
AIM@jtcir.com

Strong pace of patient enrollment supports continued advancement toward regulatory milestones

Most recent assessment of novel telomere targeting agent sequenced with a checkpoint inhibitor showed 90.5% disease control rate (DCR) in heavily pretreated NSCLC

CHICAGO, Sept. 22, 2026 (GLOBE NEWSWIRE) — MAIA Biotechnology, Inc. (NYSE American: MAIA) (“MAIA”, the “Company”), a clinical-stage biopharmaceutical company focused on developing targeted immunotherapies for cancer, today announced that enrollment has reached 65 patients in its ongoing pivotal Phase 3 trial, THIO-104, evaluating its novel telomere-targeting therapy as a third-line (3L) treatment for advanced non-small cell lung cancer (NSCLC). The THIO-104 trial currently has 38 trial sites activated in 6 foreign countries (Taiwan, Romania, Turkey, Georgia, Poland and Hungary).

“Reaching 65 randomized patients marks an important milestone in our Phase 3 trial. With additional clinical sites expected to begin enrolling patients, we remain on track to achieve our goal of more than 100 randomized patients by year-end,” said Vlad Vitoc, M.D., Founder and Chief Executive Officer of MAIA. “As we’ve stated previously, statistical assessments of the Phase 3 trial point to a very high probability of technical success for regulatory approval of ateganosine.1 We believe third-line NSCLC is an excellent market entry segment due to the substantial unmet medical need in this large immunotherapy-resistant and chemotherapy-resistant population. No current standard of care exists in this NSCLC treatment setting and competition for clinical trial patients is limited.”

In its most recent assessment, ateganosine sequenced with a checkpoint inhibitor showed 90.5% interim disease control rate (DCR) in heavily pretreated 3L NSCLC in MAIA’s ongoing phase 2 THIO-101 clinical trial. This measure contrasts with reported 25%–35% DCRs for standard third-line chemotherapy regimens.

In July 2025, the U.S. Food and Drug Administration (FDA) granted Fast Track designation for ateganosine for the treatment of NSCLC. This designation allows for more frequent FDA communication, potential rolling review, and eligibility for Accelerated Approval and Priority Review. If approved, ateganosine will hold FDA New Chemical Entity (NCE) five-year marketing exclusivity. An NCE is a small molecule drug with a novel active ingredient that hasn’t been previously approved or marketed.

About Ateganosine

Ateganosine (THIO, 6-thio-dG or 6-thio-2’-deoxyguanosine) is a first-in-class investigational telomere-targeting agent currently in clinical development to evaluate its activity in non-small cell lung cancer (NSCLC). Telomeres, along with the enzyme telomerase, play a fundamental role in the survival of cancer cells and their resistance to current therapies. The modified nucleotide 6-thio-2’-deoxyguanosine induces telomerase-dependent telomeric DNA modification, DNA damage responses, and selective cancer cell death. Ateganosine-damaged telomeric fragments accumulate in cytosolic micronuclei and activates both innate (cGAS/STING) and adaptive (T-cell) immune responses. The sequential treatment of ateganosine followed by PD-(L)1 inhibitors resulted in profound and persistent tumor regression in advanced, in vivo cancer models by induction of cancer type–specific immune memory. Ateganosine is presently developed as a second or later line of treatment for NSCLC for patients that have progressed beyond the standard-of-care regimen of existing checkpoint inhibitors.

About MAIA Biotechnology, Inc.

MAIA is a targeted therapy, immuno-oncology company focused on the development and commercialization of potential first-in-class drugs with novel mechanisms of action that are intended to meaningfully improve and extend the lives of people with cancer. Our lead program is ateganosine (THIO), a potential first-in-class cancer telomere targeting agent in clinical development for the treatment of NSCLC patients with telomerase-positive cancer cells. For more information, please visit www.maiabiotech.com.

About THIO-104 Phase 3 Clinical Trial

THIO-104 is a multicenter, open-label, randomized Phase 3 clinical trial, designed to evaluate ateganosine’s telomere-targeting anti-tumor activity when followed by PD-(L)1 inhibition in patients with advanced third-line NSCLC who previously did not respond or developed resistance to treatment regimens containing checkpoint inhibitor and/or chemotherapy and have progressed. The trial has two primary objectives: (1) to assess the clinical efficacy of ateganosine compared to investigator’s choice of chemotherapy, using median Overall Survival (OS) as the primary clinical endpoint (2) to evaluate the safety and tolerability of ateganosine in sequential combination with a checkpoint inhibitor. For more information on this Phase 3 trial, please visit ClinicalTrials.gov using the identifier NCT06908304.

Forward Looking Statements

MAIA cautions that all statements, other than statements of historical facts contained in this press release, are forward-looking statements. Forward-looking statements are subject to known and unknown risks, uncertainties, and other factors that may cause our or our industry’s actual results, levels or activity, performance or achievements to be materially different from those anticipated by such statements. The use of words such as “may,” “might,” “will,” “should,” “could,” “expect,” “plan,” “anticipate,” “believe,” “estimate,” “project,” “intend,” “future,” “potential,” or “continue,” and other similar expressions are intended to identify forward looking statements. However, the absence of these words does not mean that statements are not forward-looking. For example, all statements we make regarding (i) the initiation, timing, cost, progress and results of our preclinical and clinical studies and our research and development programs, (ii) our ability to advance product candidates into, and successfully complete, clinical studies, (iii) the timing or likelihood of regulatory filings and approvals, (iv) our ability to develop, manufacture and commercialize our product candidates and to improve the manufacturing process, (v) the rate and degree of market acceptance of our product candidates, (vi) the size and growth potential of the markets for our product candidates and our ability to serve those markets, and (vii) our expectations regarding our ability to obtain and maintain intellectual property protection for our product candidates, are forward looking. All forward-looking statements are based on current estimates, assumptions and expectations by our management that, although we believe to be reasonable, are inherently uncertain. Any forward-looking statement expressing an expectation or belief as to future events is expressed in good faith and believed to be reasonable at the time such forward-looking statement is made. However, these statements are not guarantees of future events and are subject to risks and uncertainties and other factors beyond our control that may cause actual results to differ materially from those expressed in any forward-looking statement. Any forward-looking statement speaks only as of the date on which it was made. We undertake no obligation to publicly update or revise any forward-looking statement, whether as a result of new information, future events or otherwise, except as required by law. In this release, unless the context requires otherwise, “MAIA,” “Company,” “we,” “our,” and “us” refers to MAIA Biotechnology, Inc. and its subsidiaries.

Investor Relations Contact
+1 (872) 270-3518
ir@maiabiotech.com


1 See latest MAIA investor presentation and 2026 shareholder letter at ir.maiabiotech.com/company-information/presentations.

TR-1: Standard form for notification of major holdings

NOTIFICATION OF MAJOR HOLDINGS (to be sent to the relevant issuer and to the FCA in Microsoft Word format if possible) i
 
1a. Identity of the issuer or the underlying issuer of existing shares to which voting rights are attached ii: Mothercare PLC
1b. Please indicate if the issuer is a non-UK issuer (please mark with an “X” if appropriate)
Non-UK issuer  
2. Reason for the notification (please mark the appropriate box or boxes with an “X”)
An acquisition or disposal of voting rights  

X

An acquisition or disposal of financial instruments  
An event changing the breakdown of voting rights  
Other (please specify) iii:  
3. Details of person subject to the notification obligation iv
Name Lombard Odier Asset Management (Europe) Limited
City and country of registered office (if applicable) London, United Kingdom
4. Full name of shareholder(s) (if different from 3.) v
Name Disclosure on behalf of accounts managed on a discretionary basis by Lombard Odier Investment Managers group.
City and country of registered office (if applicable)  
5. Date on which the threshold was crossed or reached vi: 18.09.2026
6. Date on which issuer notified (DD/MM/YYYY): 22.09.2026
7. Total positions of person(s) subject to the notification obligation
  % of voting rights attached to shares (total of 8. A) % of voting rights through financial instruments
(total of 8.B 1 + 8.B 2)
Total of both in % (8.A + 8.B) Total number of voting rights held in issuer (8.A + 8.B) vii
Resulting situation on the date on which threshold was crossed or reached 0.0%   0.0% 0
Position of previous notification (if
applicable)
18.93%   18.93%  

8. Notified details of the resulting situation on the date on which the threshold was crossed or reached viii
A: Voting rights attached to shares
Class/type of
shares
ISIN code (if possible)


Number of voting rights ix % of voting rights
Direct
(DTR5.1)
Indirect
(DTR5.2.1)
Direct
(DTR5.1)
Indirect
(DTR5.2.1)
GB0009067447   0   0.0%
         
         
SUBTOTAL 8. A 0 0.0%


B 1: Financial Instruments according to DTR5.3.1R (1) (a)
Type of financial instrument Expiration
date x
Exercise/
Conversion Period xi
Number of voting rights that may be acquired if the instrument is
exercised/converted.
% of voting rights


    SUBTOTAL 8. B 1    


B 2: Financial Instruments with similar economic effect according to DTR5.3.1R (1) (b)
Type of financial instrument Expiration
date x
Exercise/
Conversion Period xi
Physical or cash
Settlement xii
Number of voting rights % of voting rights


      SUBTOTAL 8.B.2    


9. Information in relation to the person subject to the notification obligation (please mark the
applicable box with an “X”)
Person subject to the notification obligation is not controlled by any natural person or legal entity and does not control any other undertaking(s) holding directly or indirectly an interest in the (underlying) issuer xiii X
Full chain of controlled undertakings through which the voting rights and/or the
financial instruments are effectively held starting with the ultimate controlling natural person or legal entity (please add additional rows as necessary) xiv
 
Name xv % of voting rights if it equals or is higher than the notifiable threshold % of voting rights through financial instruments if it equals or is higher than the notifiable threshold Total of both if it equals or is higher than the notifiable threshold


10. In case of proxy voting, please identify:
Name of the proxy holder  
The number and % of voting rights held  
The date until which the voting rights will be held  
 
11. Additional information xvi
 

Place of completion London, United Kingdom
Date of completion 22.09.2026

Collaboration builds on Itron’s proven security-first platform and integrates Crytica’s Rapid Detection, Alert and Isolation technology to help utilities prepare for emerging cyber and operational risks at the grid edge

LIBERTY LAKE, Wash., Sept. 22, 2026 (GLOBE NEWSWIRE) — Itron, Inc. (NASDAQ: ITRI), the intelligent infrastructure provider for modern energy and water management, and Crytica Security announced a strategic technology collaboration to advance rapid cyber threat detection and operational visibility across its intelligent grid-edge infrastructure.

Cybersecurity threats to critical infrastructure are ever-present and evolving rapidly. As utilities modernize their infrastructure, deploy more distributed intelligence at the grid edge, integrate distributed energy resources and prepare for new AI-enabled threat vectors, security must adapt. Utilities require solutions that build on proven protection while adapting faster to new risks emerging across distributed, connected and intelligent networks.

The collaboration is designed to integrate Crytica’s patented Rapid Detection, Alert and Isolation technology, RDAi™, into Itron’s Grid Edge Intelligence portfolio, adding continuous integrity monitoring and actionable alerts closer to where grid activity occurs. The combination of Itron’s secure, utility-grade platform with Crytica’s deterministic detection technology, provides utilities with an additional layer of cybersecurity visibility across IT and OT environments and intelligent grid edge devices, meter-hosted applications and edge workloads as the security landscape evolves.

Itron’s Grid Edge Intelligence portfolio is built on a strong security foundation that includes layered device, network, communications and data protection. The integration of Crytica’s RDAi technology is intended to extend that foundation with active, embedded monitoring that can detect unauthorized changes to devices without negatively affecting device performance. This added layer helps utilities identify potential compromise earlier, accelerate investigation and response, and strengthen resilience across critical field assets.

“Utilities are moving intelligence closer to the edge of the grid, and cybersecurity must move with it,” said Don Reeves, Itron’s senior vice president of Outcomes. “Itron has long helped utilities deploy secure, reliable and resilient infrastructure at scale. Our collaboration with Crytica builds on that foundation by adding rapid, embedded threat detection designed for the next generation of grid-edge applications. Together, we are helping utilities prepare for what is coming next while continuing to protect the essential services communities rely on every day.”

Unlike traditional cybersecurity approaches that focus primarily on centralized systems or enterprise endpoints, the Itron-Crytica collaboration is designed to bring monitoring directly into resource-constrained utility field devices and the applications running on them. Crytica’s lightweight probe continuously monitors and delivers high-fidelity alerts when unauthorized changes are detected. This approach is designed to complement existing cybersecurity platforms by extending visibility into operational technology environments where traditional endpoint tools may not operate.

This collaboration aims to advance embedded grid-edge threat detection by helping utilities monitor both the meter and meter-hosted applications through a common integrity monitoring framework, a capability not available to utilities today.

The collaboration also supports broader operational awareness. In addition to cybersecurity monitoring, the integration is expected to provide performance-related alerts that help utilities identify conditions that may affect device health, reliability and availability. These capabilities can help utilities improve situational awareness, prioritize investigation and support more resilient operations as AMI networks evolve into intelligent grid-edge platforms.

“Crytica’s RDAi technology was purpose-built for demanding, resource-constrained operational environments,” said C. Lloyd Mahaffey, Executive Chairman and Co-Founder, Crytica Security. “By working with Itron, we can extend rapid detection and actionable alerts into utility grid-edge infrastructure at scale, helping utilities see potential issues earlier and respond with greater confidence.”

The collaboration reinforces Itron’s strategy to help utilities transition from traditional AMI to Grid Edge Intelligence by combining distributed intelligence, intelligent connectivity, advanced analytics and security-centered innovation. As utilities prepare for more dynamic grid operations, the ability to detect, alert and respond closer to the edge will become increasingly important to maintaining reliability, resilience and trust.

Customer benefits

  • Builds on a proven security foundation: Extends Itron’s secure, utility-grade platform with an additional layer of embedded threat detection.
  • Strengthens grid-edge visibility: Helps utilities monitor intelligent devices, meter-hosted applications and edge workloads closer to where grid activity occurs.
  • Enables faster detection and response: Uses Crytica’s RDAi technology to identify unauthorized instruction-set changes within seconds and generate actionable alerts.
  • Supports operational resilience: Adds visibility into device health and performance conditions that may affect reliability, availability or service delivery.
  • Prepares utilities for emerging threats: Helps utilities adapt to an evolving threat landscape, including risks associated with increasing connectivity, distributed intelligence and AI-enabled cyber activity.

About Itron
Itron is transforming how the world manages energy, water and city services. Our trusted intelligent infrastructure solutions help utilities and cities improve efficiency, build resilience and deliver safe, reliable and affordable service. With edge intelligence, we connect people, data insights and devices so communities can better manage the essential resources they rely on to live and thrive. Join us as we create a more resourceful world: www.itron.com

About Crytica Security
Crytica Security’s patented Rapid Detection, Alert (RDAi™), and Isolation technology embeds a lightweight, under 100 Kilobyte, Probe directly within operational technology, IoT, embedded systems and other cyber-physical environments to continuously monitor system integrity and detect unauthorized changes to instruction sets within seconds. Designed to complement existing cybersecurity platforms, RDAi extends visibility into devices where traditional tools may not operate, delivering deterministic detection and actionable alerts that enable faster security response for mission-critical operations. See how RDAi can help you “detect and protect” by visiting www.cryticasecurity.com.

Media Contacts:
Itron, Inc.
Alison Mallahan
Senior Manager, Corporate Communications
509-891-3802
PR@Itron.com

Paul Vincent
Vice President, Investor Relations
512-560-1172
Investors@itron.com

Crytica Security, Inc.
Jake Blanchard, VP, OEM Sales
Email: jake@cryticasecurity.com
Mobile: (208) 559-3399

MINNEAPOLIS, Sept. 22, 2026 (GLOBE NEWSWIRE) — OneMedNet Corporation (Nasdaq: ONMD) (“OneMedNet,” the “Company,” “we,” “us,” or “our”), a leading provider of regulatory decision-grade, AI-driven Real-World Data (RWD), today announced a strategic partnership with Atropos Health, the creator of real-world evidence for clinical decision-making under a multi-year agreement. The partnership adds OneMedNet’s regulatory decision-grade data to Atropos’ platform and customer offerings under a fee arrangement adding to our Annual Recurring Revenue (ARR).

The partnership embeds OneMedNet’s data directly into Atropos Health’s evidence-generation platform, giving Atropos customers access to more multi-modal data drawn from our network of over 90 million patient journeys. Our multi-modal RWD brings clinical depth that elevates evidence quality well beyond what claims or Electronic Health Record (EHR) datasets alone can deliver, positioning our RWD as differentiated, high-value data within the partnership.

Under the partnership, Atropos’ customers – health systems, life sciences organizations, and researchers – will be able to purchase additional OneMedNet multi-modal data from OneMedNet through the Atropos platform. The collaboration is structured as a 3-year subscription-based license purchased by Atropos, providing seamless, ongoing data access and data delivery.

Solving the Evidence Gap Together

Most Real-World Evidence today lacks the clinical richness of imaging and multimodal data. Atropos Health solves for speed and rigor by adding imaging to its tools, providing more real-world evidence than any other platform. OneMedNet supplies the regulatory-grade depth and breadth both companies believe is essential for truly credible, decision-grade evidence. The subscription partnership unites these strengths, making OneMedNet’s differentiated data a core, recurring component of Atropos’ platform workflows.

“This partnership perfectly exemplifies our strategy: transforming our unique, Direct-from-Source, regulatory decision-grade data into long-term subscription collaborations that put our data directly into our partners’ hands removing the friction that has traditionally slowed access to high-quality Real-World Data,” said Aaron Green, President & CEO of OneMedNet. “Atropos sets the standard for rapid, personalized evidence, and we strengthen the foundational data beneath it so their customers can move from question to evidence without the usual barriers.”

“High-quality evidence starts with fit-for-purpose data, and multimodal depth is now table stakes,” said Dr. Brigham Hyde, CEO and Co-Founder of Atropos Health. “Our partnership with OneMedNet brings regulatory-grade imaging and multimodal richness into our platform, enabling richer insights for our clients as we work together to close the evidence gap.”

The collaboration reflects surging demand for our multimodal, regulatory-grade RWD and further validates OneMedNet’s focus on converting differentiated data assets into long-term, partnership-driven value.

About OneMedNet Corporation
OneMedNet Corporation is revolutionizing Real-World Data (RWD) through its iRWD™ platform, delivering regulatory decision-grade, AI-ready datasets that include de-identified medical imaging alongside comprehensive clinical records. With a network spanning more than 2,300 sites and encompassing over 90 million patient journeys and 270 million studies, OneMedNet serves life sciences companies, medical device manufacturers, AI developers, and other innovators seeking high-quality, compliant healthcare data. The Company’s platform supports applications ranging from drug development and regulatory submissions to foundational AI model training. Learn more at www.onemednet.com.

Forward-Looking Statements

This press release contains forward-looking statements within the meaning of Section 27A of the Securities Act of 1933, as amended, and Section 21E of the Securities Exchange Act of 1934, as amended. All statements other than statements of historical facts contained in this press release are forward-looking statements. These statements include, but are not limited to, the benefits of the Atropos Health partnership, statements regarding demand for our products and services, the composition and durability of our customer base, the speed of order fulfillment, the acceptance of our delivered data sets allowing for the conversion of bookings to revenue, our ability to achieve our operational strategies, our expected growth in the business, and statements generally about our products, plans and strategies.

Forward-looking statements are based on information available at the time those statements are made and were based on current expectations as well as the beliefs and assumptions of management as of that time with respect to future events. These statements are subject to risks and uncertainties, many of which involve factors or circumstances that are beyond our control. These risks and uncertainties include, but are not limited to: our ability to change the direction of OneMedNet; our ability to keep pace with new technology and changing market needs; the competitive environment of our business; the timing of customer acceptance and data delivery; and the other risks described in our most recent Annual Report on Form 10-K and our subsequent filings with the Securities and Exchange Commission. Except as required by law, we do not undertake any obligation to publicly update or revise any forward-looking statement, whether as a result of new information, future developments, or otherwise.

OneMedNet Contacts:
Michael Wong, VP Marketing
Email: michael.wong@onemednet.com
SOURCE: ONEMEDNET CORPORATION

Palisade Bio, Inc.

Phase 2 ASCENTRA-CD clinical trial in patients with moderate to severe Crohn’s disease (CD)

ASCENTRA-CD expected to enroll approximately 60 patients with moderate to severe Crohn’s disease

First patient dosing targeted for Q1 2027; primary efficacy readout (Week 12 endoscopic response) targeted for Q1 2028

Phase 2 program builds on clinical experience in fibrostenotic Crohn’s disease demonstrating targeted tissue exposure, PDE4 target engagement and encouraging endoscopic activity

Denver, CO, Sept. 22, 2026 (GLOBE NEWSWIRE) — Palisade Bio, Inc. (Nasdaq: PALI) (“Palisade” or the “Company”), a clinical-stage biopharmaceutical company developing next-generation prodrugs for patients living with inflammatory and fibrotic diseases, today announced that the U.S. Food and Drug Administration (FDA) has cleared the Company’s Investigational New Drug (IND) application for PALI-2108, enabling initiation of the Phase 2 ASCENTRA-CD clinical trial in patients with moderate to severe Crohn’s disease (CD).

“FDA clearance of the ASCENTRA-CD IND marks another significant milestone for PALI-2108 and meaningfully expands our Phase 2 development program across inflammatory bowel disease,” said Mitch Jones, M.D., Ph.D., President and Chief Medical Officer of Palisade Bio. “Crohn’s disease remains a progressive condition where many patients experience inadequate response or loss of response to existing therapies. We believe PALI-2108’s differentiated prodrug design, once-daily oral dosing and targeted activation within the ileum and colon provide a compelling foundation for development in this population. Importantly, we enter Phase 2 with direct clinical experience demonstrating favorable tolerability, targeted tissue exposure, PDE4 target engagement and encouraging early biological and endoscopic activity.”

ASCENTRA-CD Phase 2 Trial Design

ASCENTRA-CD is a multicenter, open-label, single-arm Phase 2 study designed to evaluate the efficacy, safety, pharmacokinetics and pharmacodynamics of PALI-2108 in approximately 60 adults with moderate to severe Crohn’s disease. Patients will undergo a 14-day dose titration to the target dose of PALI-2108 30 mg orally once daily. The study includes a 12-week induction phase followed by a 36-week maintenance phase.

The primary endpoint is endoscopic response at Week 12, defined as at least a 50% reduction from baseline in the Simple Endoscopic Score for Crohn’s Disease (SES-CD), based on blinded central endoscopic review. Key secondary endpoints at Week 12 include clinical remission, endoscopic remission, PRO-2 remission (a patient-reported outcome of abdominal pain and stool frequency), clinical response and combined clinical remission and endoscopic response, with additional maintenance assessments through Week 48.

The Company anticipates first patient dosing in Q1 2027, and a primary efficacy readout in Q1 2028.

PALI-2108 in Crohn’s Disease

Crohn’s disease is a chronic, progressive inflammatory condition in which persistent or recurrent inflammation can lead to irreversible tissue remodeling, fibrosis and bowel disability.

PALI-2108 is designed to address these challenges through an oral, locally activated prodrug approach. Following oral administration, the drug is activated in the terminal ileum and colon, generating high local intestinal exposure while also supporting sustained systemic exposure and a differentiated pharmacokinetic profile. PALI-2108 has already been evaluated in patients with fibrostenotic Crohn’s disease, providing direct clinical experience with ileal and colonic tissue exposure, PDE4 target engagement and endoscopic activity. ASCENTRA-CD is designed to extend those findings into a broader moderate to severe Crohn’s disease population using centrally read endoscopic response at Week 12 as the primary endpoint.

With Phase 2 development now advancing in both ulcerative colitis and Crohn’s disease, Palisade is advancing a broad clinical development program for PALI-2108 across the two major forms of inflammatory bowel disease. Together, ASCENTRA-UC in ulcerative colitis and ASCENTRA-CD in Crohn’s disease are designed to generate a broader clinical dataset around the efficacy, safety and pharmacology of PALI-2108.

About PALI-2108

PALI-2108 is a once-daily oral PDE4 inhibitor prodrug designed for targeted activation in the terminal ileum and colon. Across Phase 1 studies in healthy volunteers and patients with ulcerative colitis and fibrostenotic Crohn’s disease, PALI-2108 demonstrated favorable safety and tolerability, differentiated pharmacokinetics and sustained PDE4 inhibition above IC90, supporting advancement into Phase 2 development.

About Palisade Bio

Palisade Bio, Inc. (Nasdaq: PALI) is a clinical-stage biopharmaceutical company developing prodrugs for patients living with inflammatory and fibrotic diseases.

The Company’s lead clinical product candidate, PALI-2108, is being advanced into Phase 2 clinical trials as a treatment for patients living with inflammatory bowel disease (IBD), including ulcerative colitis (UC) and Crohn’s disease (CD). Despite the availability of multiple biologic and small-molecule therapies, many patients with UC and CD do not achieve durable remission, lose response over time, or discontinue treatment because of safety or tolerability limitations. The Company believes PALI-2108 has the potential to address these limitations through once-daily oral dosing, targeted lower-intestinal bioactivation and broad PDE4-mediated anti-inflammatory activity.

For more information, please go to www.palisadebio.com.

Forward Looking Statements

Any statements contained in this communication that are not statements of historical fact may be deemed to be forward-looking statements for purposes of the safe harbor provisions of the Private Securities Litigation Reform Act of 1995. Forward-looking statements include but are not limited to: implied or express statements regarding the pharmacological properties, safety, tolerability, clinical response and efficacy, and therapeutic potential of PALI-2108, dosing levels and timing of key PALI-2108 development milestones such as regulatory submissions and approvals, trial enrollments and commencements and efficacy readouts, and the Company’s expected cash runway. These forward-looking statements are based on the Company’s current expectations. Forward-looking statements involve risks and uncertainties. Important factors that could cause actual results to differ materially from those reflected in the Company’s forward-looking statements include, among others, the timing of enrollment, commencement and completion of the Company’s clinical trials; the Company’s reliance on PALI-2108, and its early stage of clinical development; the risk that prior results, such as signals of safety, clinical response and efficacy, dosing or durability of effect, observed from preclinical or clinical trials with a limited number of patients, will not be replicated or will not continue in ongoing or future studies or clinical trials involving the Company’s product candidates in clinical trials focused on the same or different indications; and other factors that are described in the “Risk Factors” and “Management’s Discussion and Analysis of Financial Condition and Results of Operations” sections of the Company’s Annual Report on Form 10-K for the fiscal year ended December 31, 2025, filed with the Securities and Exchange Commission (“SEC”) on March 20, 2026, and the Quarterly Reports on Form 10-Q or other SEC filings that are filed thereafter. Investors are cautioned not to put undue reliance on these forward-looking statements. These forward-looking statements speak only as of the date hereof, and the Company expressly disclaims any obligation or undertaking to release publicly any updates or revisions to any forward-looking statements contained herein to reflect any change in the Company’s expectations with regard thereto or any change in events, conditions or circumstances on which any such statements are based, except as required by law.

Investor Relations Contact

JTC Team, LLC
Jenene Thomas
908-824-0775
PALI@jtcir.com

Attachment

First patient marks Polyrizon’s transition into a clinical-stage company.

The U.S clinical study is designed to evaluate the efficacy, safety and tolerability of the NASARIX (PL-14) Allergy Blocker in patients with seasonal allergic rhinitis

Raanana, Israel, Sept. 22, 2026 (GLOBE NEWSWIRE) — Polyrizon Ltd. (Nasdaq: PLRZ) (the “Company” or “Polyrizon”), a clinical- stage biotech company specializing in the development of innovative intranasal hydrogels, today announced the enrollment of the first patient in its U.S. clinical study to evaluate NASARIX (PL-14), the Company’s investigational Allergy Blocker, in patients with seasonal allergic rhinitis (SAR).

The enrollment of the first patient represents a significant milestone for Polyrizon and marks the Company’s transition into the clinical stage of development.

“This is a defining milestone for Polyrizon,” said Tomer Izraeli, Chief Executive Officer of Polyrizon. “The enrollment of the first patient in our U.S. study represents the culmination of extensive product development, preclinical evaluation, manufacturing and regulatory preparation, and moves NASARIX into clinical evaluation in patients for the first time. We believe this study represents an important step toward our goal of developing a differentiated, drug-free approach for people suffering from seasonal allergies.”

NASARIX is a non-pharmacological, drug-free, mucoadhesive hydrogel nasal spray designed to form a physical barrier within the nasal cavity. The barrier is intended to reduce contact between inhaled airborne allergens and the nasal mucosa, thereby helping alleviate symptoms associated with allergic rhinitis.

The prospective, multicenter, randomized, single-blind, two-arm study is expected to randomize 120 patients at 5 clinical sites in the United States. Participants will be randomized 1:1 to receive either NASARIX Allergy Blocker or a saline nasal spray comparator.

The study is designed to evaluate the efficacy, safety and tolerability of NASARIX in adults with seasonal allergic rhinitis. The primary efficacy endpoint will assess the difference in mean daily reflective Total Nasal Symptom Score (rTNSS) between PL-14 and the saline comparator over the first 14 treatment days. Safety assessments will include the frequency and severity of treatment-emergent adverse events as well as assessments of local and overall tolerability.

“Moving the NASARIX Allergy Blocker into a first-in-human clinical study is an important step in the development of our Capture and Contain™ technology from concept through product development and preclinical evaluation,” said Dr. Tidhar Turgeman, Chief Technology Officer of Polyrizon. “The study has been designed to generate data regarding efficacy and safety under real-world seasonal allergen exposure. We look forward to advancing enrollment across our U.S. clinical sites and evaluating NASARIX’s potential as a drug-free option for patients with seasonal allergic rhinitis.”

About NASARIXTM

NASARIXTM is Polyrizon’s proprietary nasal spray designed to create a protective barrier within the nasal cavity, helping to prevent contact between inhaled allergens and the nasal mucosa. The product is being developed as a non-drug approach for the management of seasonal allergic rhinitis.

About Polyrizon

Polyrizon is a clinical- stage biotech company specializing in the development of innovative medical device hydrogels delivered in the form of nasal sprays, which form a thin hydrogel-based shield containment barrier in the nasal cavity that can provide a barrier against viruses and allergens from contacting the nasal epithelial tissue. Polyrizon’s proprietary Capture and Contain TM, or C&C, hydrogel technology, comprised of a mixture of naturally occurring building blocks, is delivered in the form of nasal sprays, and potentially functions as a “biological mask” with a thin shield containment barrier in the nasal cavity. Polyrizon is further developing certain aspects of its C&C hydrogel technology, such as the bioadhesion and prolonged retention at the nasal deposition site for intranasal delivery of drugs. Polyrizon refers to its additional technology, which is in an earlier stage of pre-clinical development, that is focused on nasal delivery of active pharmaceutical ingredients, or APIs, as Trap and Target, or T&T. For more information, please visit https://polyrizon-biotech.com.

Forward Looking Statements

This press release contains “forward-looking statements” within the meaning of the Private Securities Litigation Reform Act of 1995 and other securities laws. Words such as “expects,” “anticipates,” “intends,” “plans,” “believes,” “seeks,” “estimates” and similar expressions or variations of such words are intended to identify forward-looking statements. For example, the Company is using forward-looking statements when it discusses the expected conduct, timing and progress of the NASARIX (PL-14) clinical study, including patient enrollment, the anticipated number of patients and clinical sites, the study’s evaluation of efficacy, safety and tolerability, the generation of clinical data, and NASARIX’s potential as a drug-free option for patients with seasonal allergic rhinitis. Forward-looking statements are not historical facts, and are based upon management’s current expectations, beliefs and projections, many of which, by their nature, are inherently uncertain. Such expectations, beliefs and projections are expressed in good faith. However, there can be no assurance that management’s expectations, beliefs and projections will be achieved, and actual results may differ materially from what is expressed in or indicated by the forward-looking statements. Forward-looking statements are subject to risks and uncertainties that could cause actual performance or results to differ materially from those expressed in the forward-looking statements. For a more detailed description of the risks and uncertainties affecting the Company, reference is made to the Company’s reports filed from time to time with the Securities and Exchange Commission (“SEC”), including, but not limited to, the risks detailed in the Company’s Annual Report on Form 20-F for the fiscal year ended December 31, 2025 and subsequent filings with the SEC. Forward-looking statements speak only as of the date the statements are made. The Company assumes no obligation to update forward-looking statements to reflect actual results, subsequent events or circumstances, changes in assumptions or changes in other factors affecting forward-looking information except to the extent required by applicable securities laws. If the Company does update one or more forward-looking statements, no inference should be drawn that the Company will make additional updates with respect thereto or with respect to other forward-looking statements. References and links to websites have been provided as a convenience, and the information contained on such websites is not incorporated by reference into this press release. Polyrizon is not responsible for the contents of third-party websites.

Contacts:

Michal Efraty
Investor Relations
IR@polyrizon-biotech.com

Colorado PPE

Colorado PPE
Colorado PPE

Colorado PPE

Colorado PPE
Colorado PPE

Colorado PPE

Colorado PPE
Colorado PPE

Colorado PPE

Colorado PPE
Colorado PPE

Colorado PPE

Colorado PPE
Colorado PPE

Colorado PPE

Colorado PPE
Colorado PPE

Expands Access to Comprehensive Firefighter PPE Care with the Opening of its Colorado Facility

New Facility Provides Fire Departments with Cleaning, Inspection, Repair, Rental PPE, NFPA 1850 Training and Department Cleaning Equipment Under One Roof

HUNTSVILLE, Ala., Sept. 22, 2026 (GLOBE NEWSWIRE) — Lakeland Industries, Inc. (“Lakeland Fire + Safety” or “Lakeland”) (NASDAQ: LAKE) announced today the expansion of its network of independent service providers for comprehensive firefighter PPE care with the opening of its new Centennial, Colorado facility, Colorado PPE. Colorado PPE combines responsive department-focused service with the resources and capabilities of Lakeland Fire + Safety to deliver comprehensive PPE care, rental and training solutions.

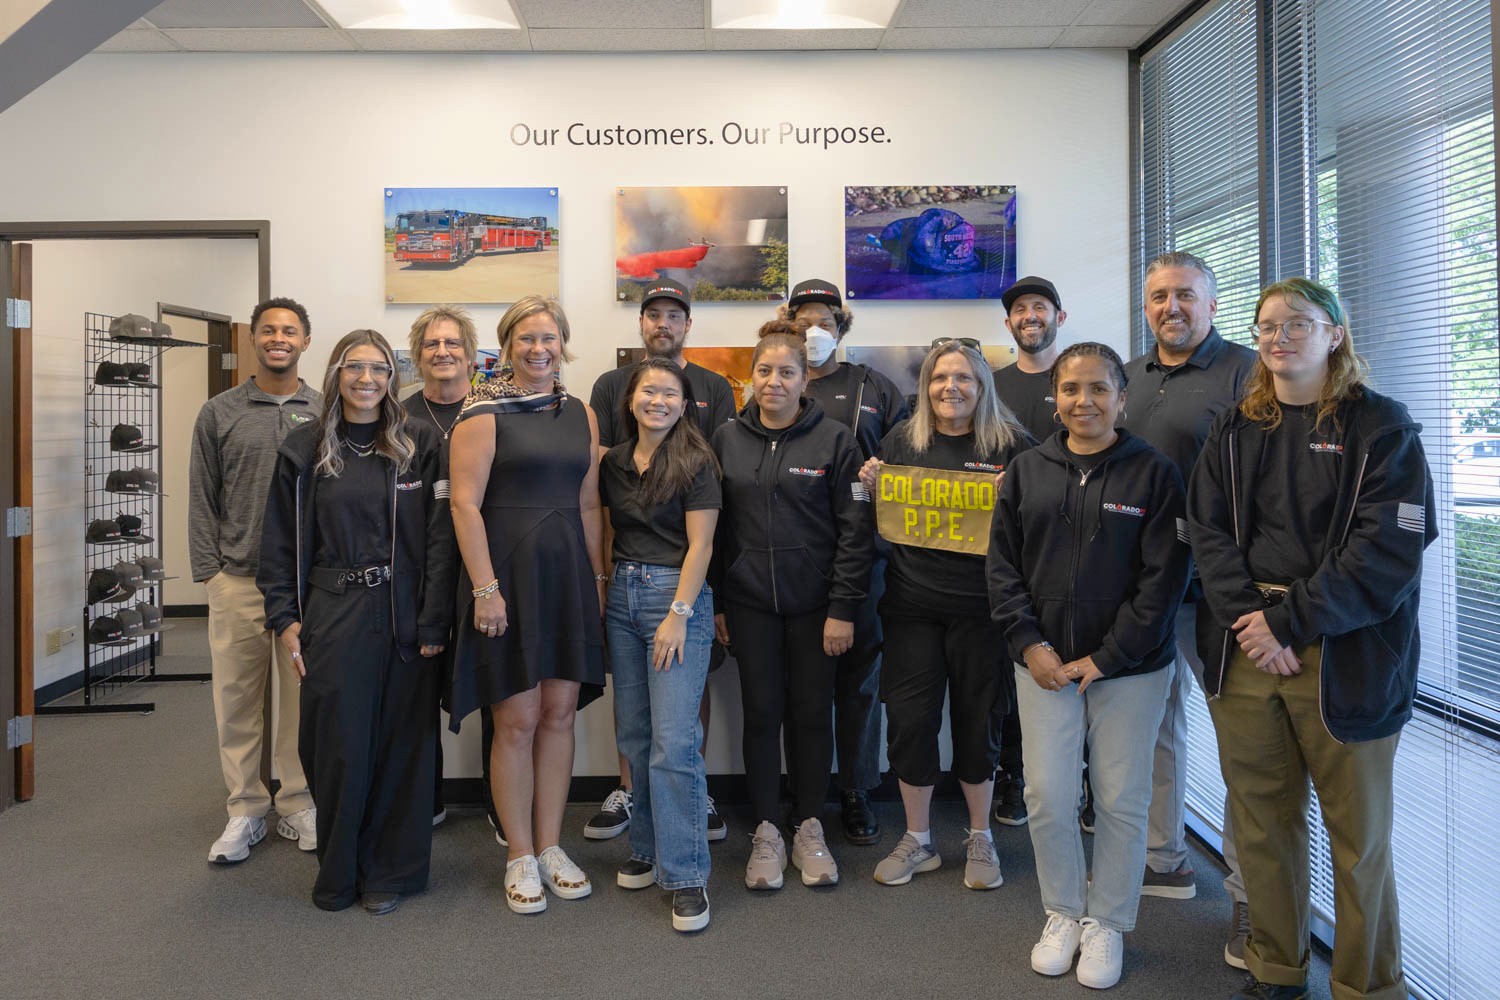
LAKE_Photos_COPPE-LS-Edits_09.2026-141

Colorado PPE operates as an independent service provider, offering fire departments a more personal approach while bringing the resources and capabilities of a larger organization to the relationship. The facility represents a broader approach to firefighter PPE management—one that goes beyond compliance to focus on the people who rely on their gear every time they answer a call.

“We believe in maintaining a higher level of service for our fire departments,” said Mike Glaze, President of California PPE. “Our goal is to ensure our firefighters and departments are valued and their gear is in the best condition it can be in to protect them when it leaves our facility and enters their service.”

NFPA 1850, is the National Fire Protection Association standard that establishes requirements for the selection, care, maintenance, inspection, cleaning, repair, retirement, and management of protective ensembles and clothing used by firefighters and emergency responders. Under this standard, it establishes a minimum performance benchmark of removing 50% of heavy metals, polycyclic aromatic hydrocarbons (PAHs), and volatile organic compounds (VOCs) from contaminated PPE. Based on test results from UL, a third-party testing agency, our cleaning process achieved average cleaning efficiencies of 88% removal of VOCs and 89% removal of heavy metals—exceeding the minimum benchmark and raising the bar for PPE cleaning in the industry.

The new facility provides comprehensive PPE services designed to support firefighters throughout the lifecycle of their gear, including cleaning, inspection and repair. Rental PPE is also available to help departments maintain readiness when gear is temporarily out of service or when additional equipment is needed for training and other operational requirements. Colorado PPE will also provide NFPA 1850 training and station cleaning equipment such as extractors to help departments better understand and implement effective PPE care programs.

“Every department has different needs, different resources and different challenges. Our job is to understand those needs and help build a PPE care program that works for them,” said Misty Johnson, Operations Manager at Colorado PPE. “We clean, inspect and repair PPE with purpose—going beyond the minimum to help protect those who protect us because when it comes to protecting firefighters, meeting the minimum should be the starting point, not the finish line.”

Jim Jenkins, President and Chief Executive Officer of Lakeland, added, “Colorado PPE is committed to helping departments care for their gear with purpose and looks forward to serving the Colorado fire community. PPE care is ultimately about more than equipment, standards or processes. It is about the people wearing the gear. The firefighters who run toward emergencies trust their PPE to provide protection in some of the most demanding environments imaginable and Colorado PPE believes the care that gear receives should reflect the importance of that responsibility.”

About Lakeland Fire + Safety

Lakeland Fire + Safety manufactures and sells a comprehensive line of fire services and industrial protective clothing and accessories for the industrial and first responder markets. In addition, we provide decontamination, repair and rental services that complement our fire services portfolio. Our products are sold globally by our in-house sales teams, our customer service group, and authorized independent sales representatives to a strategic global network of selective fire and industrial distributors and wholesale partners. Our authorized distributors supply end users across various industries, including integrated oil, chemical/petrochemical, automobile, transportation, steel, glass, construction, smelting, cleanroom, janitorial, pharmaceutical, and high-tech electronics manufacturers, as well as scientific, medical laboratories, and the utilities industry. In addition, we supply federal, state and local governmental agencies and departments, including fire and law enforcement, airport crash rescue units, the Department of Defense, the Department of Homeland Security and the Centers for Disease Control. Internationally, we sell to a mix of end-users directly and to industrial distributors, depending on the particular country and market. In addition to the United States, sales are made into more than 50 foreign countries, the majority of which were into China, the European Economic Community (“EEC”), Canada, Chile, Argentina, Commonwealth of Independent States (“CIS”) Region, Colombia, Mexico, Ecuador, India, Uruguay, Middle East, Southeast Asia, Australia, Hong Kong and New Zealand.

For more information about Lakeland, please visit the Company’s website at www.lakeland.com.

About Colorado PPE

Located in Centennial, Colorado, Colorado PPE provides comprehensive PPE care and resources for fire departments, including cleaning, inspection, repair, rental PPE, NFPA 1850 training and equipment, and advanced decontamination solutions.

Colorado PPE combines the responsive, department-focused approach of an independent service provider with the resources, technical expertise and broader capabilities of Lakeland Fire + Safety. This combination allows Colorado PPE to deliver personal service at the local level while continuing to invest in technologies and capabilities designed to improve firefighter safety and PPE care. For more information visit https://coloradoppe.com/.

Colorado PPE
6675 S. Kenton St., Suite 113
Centennial, CO 80111

Hours: Monday–Friday, 8:00 a.m.-3:30 p.m. MT
Email: customerservice@coloradoppe.com

“Safe Harbor” Statement under the Private Securities Litigation Reform Act of 1995

This press release contains estimates, predictions, opinions, goals and other “forward-looking statements” as that phrase is defined in the Private Securities Litigation Reform Act of 1995. Such statements include, without limitation, references to the Company’s predictions or expectations of future business or financial performance as well as its goals and objectives for future operations, financial and business trends, business prospects, and management’s expectations for earnings, revenues, expenses, inventory levels, capital levels, liquidity levels, or other future financial or business performance, strategies or expectations, including without limitation our plans to deliver comprehensive care, rental and training solutions. All statements, other than statements of historical facts, which address Lakeland’s expectations of sources or uses for capital, or which express the Company’s expectation for the future with respect to financial performance or operating strategies, can be identified as forward-looking statements. Forward-looking statements involve risks, uncertainties and assumptions as described from time to time in press releases and Forms 8-K, registration statements, quarterly and annual reports and other reports and filings filed with the Securities and Exchange Commission or made by management. As a result, there can be no assurance that Lakeland’s future results will not be materially different from those described herein as “believed,” “projected,” “planned,” “intended,” “anticipated,” “can,” “estimated” or “expected,” or other words which reflect the current view of the Company with respect to future events. We caution readers that these forward-looking statements speak only as of the date hereof. The Company hereby expressly disclaims any obligation or undertaking to release publicly any updates or revisions to any such statements to reflect any change in the Company’s expectations or any change in events, conditions or circumstances on which such statement is based, except as may be required by law.

Investor Relations
Chris Tyson
Executive Vice President
MZ Group – MZ North America
949-491-8235
LAKE@mzgroup.us 
www.mzgroup.us

Photos accompanying this announcement are available at:

https://www.globenewswire.com/NewsRoom/AttachmentNg/1466df9b-0c9b-4b3c-a9ff-e4d7ba3b6eeb

https://www.globenewswire.com/NewsRoom/AttachmentNg/9ea55120-6143-4807-9908-cd5baf7bf773

https://www.globenewswire.com/NewsRoom/AttachmentNg/ae36e0fd-d833-4128-9654-cf7c4f179bac

https://www.globenewswire.com/NewsRoom/AttachmentNg/225c9d58-004e-40c7-b0fb-b9c41d3d91d6

https://www.globenewswire.com/NewsRoom/AttachmentNg/1eddf22f-adef-42a4-b74e-81a0a27d0cb3

https://www.globenewswire.com/NewsRoom/AttachmentNg/156dd466-2fdd-4937-b313-66a65fa7ec75

https://www.globenewswire.com/NewsRoom/AttachmentNg/913d3647-aadc-41cb-923b-ff7c5e36cb7a 

Approved Exploration Plan of Operations and 2026 Annual Operating Plan Clear the Way for Drilling; Company Targeting Up to 15,000 Feet of Additional Diamond Core Drilling This Season

BOISE, Idaho, Sept. 22, 2026 (GLOBE NEWSWIRE) — Idaho Copper Corporation (NYSE American: COPR) (“Idaho Copper” or the “Company”), a critical minerals developer advancing the flagship CuMo copper-molybdenum-silver project in Idaho, today announced that the U.S. Forest Service, Boise National Forest (“USFS”) sent a letter approving the Company’s Exploration Plan of Operations (“PoO”) and its 2026 annual operating plan for exploration at the CuMo project in Boise County, Idaho, clearing the way for drilling and roadbuilding and maintenance activities to commence.

On September 9, 2026, the USFS accepted the Company’s signed Forest Service Evaluation form and incorporated all required changes, modifications and mitigations into the approved PoO, and has accepted the Company’s reclamation surety bond. The letter further notes that the Company has provided documentation showing that all other permits necessary for the project are in place and in good standing. The USFS also approved the Company’s 2026 annual operating plan, which was reviewed and field verified by agency staff. The approved PoO is effective for six years from the first day of operations, inclusive of final reclamation.

The approval follows the Decision Notice and Finding of No Significant Impact issued by the USFS on March 14, 2025 under an Environmental Assessment, which completed the federal environmental review of the CuMo exploration plan. The September 9 letter satisfies the remaining administrative conditions to that decision and authorizes the Company to begin work.

Management Commentary

“Approval of our Exploration Plan of Operations and our 2026 annual operating plan is the culmination of a decade of work with the Forest Service, and it is the milestone that should turn CuMo from a concept into an active drill program,” said Andrew Brodkey, Chief Executive Officer of Idaho Copper. “We have an approved plan, an accepted reclamation bond, and the other permits this project requires. In a country where federal permitting is often the single largest obstacle impeding advancement of a critical minerals deposit, that is a meaningful achievement.”

“Our priority for the balance of this season is infill drilling that tightens confidence in the existing deposit, and we plan to carry that work into a substantially larger campaign in 2027 aimed at completing the drilling needed for a Prefeasibility Study. Every foot we drill from here will be directed at advancing CuMo toward a development decision, and at doing so under the environmental commitments we have made to the Forest Service and to the communities around this project,” concluded Brodkey.

About Idaho Copper Corporation

Idaho Copper Corporation (NYSE American: COPR) is a critical minerals developer focused on exploring and developing the CuMo copper-molybdenum-silver-tungsten-rhenium project located in Boise County, Idaho. The CuMo project is one of the largest undeveloped copper deposits in the U.S., which management believes is among the largest undeveloped primary molybdenum deposits in the world, which also contains significant amounts of silver, rhenium, and tungsten — all considered critical or of strategic importance. The project comprises approximately 2,640 acres and consists of 126 federal unpatented lode mining claims and 6 patented mining claims. To learn more, please visit www.idaho-copper.com.

Safe Harbor Statement

With the exception of historical information contained in this press release, content herein may contain “forward-looking statements” that are made pursuant to the Safe Harbor Provisions of the U.S. Private Securities Litigation Reform Act of 1995. Forward-looking statements are generally identified by using words such as “anticipate,” “believe,” “plan,” “expect,” “intend,” “will,” and similar expressions, but these words are not the exclusive means of identifying forward-looking statements. Forward-looking statements in this release include statements regarding the Company’s 2026 and 2027 exploration and drilling programs, the expected footage, scope and timing thereof, and the planned Prefeasibility Study and the anticipated timing thereof. These statements are based on management’s current expectations and are subject to uncertainty and changes in circumstances. Investors are cautioned that forward-looking statements involve risks and uncertainties that could cause actual results to differ materially from the statements made, including risks related to metal price volatility, mining and processing performance, drill rig and contractor availability, weather and the length of the permitted operating season, permitting, and the availability of financing for and development of the CuMo project. Important factors that could cause our actual results and financial condition to differ materially from those indicated in the forward-looking statements are discussed or identified in our filings with the Securities and Exchange Commission (the “SEC”), including the risk factors contained in our most recent Annual Report on Form 10-K, Quarterly Reports on Form 10-Q, and registration statements. In addition, this press release contains time-sensitive information that reflects management’s best analysis only as of the date of this press release. Idaho Copper does not undertake any obligation to publicly update or revise any forward-looking statements to reflect future events, information or circumstances that arise after the date of this release.

Investor Relations Contact:
Lucas A. Zimmerman
Managing Director
MZ Group – MZ North America
(262) 357-2918
COPR@mzgroup.us
www.mzgroup.us

Privacy Overview

This website uses cookies so that we can provide you with the best user experience possible. Cookie information is stored in your browser and performs functions such as recognising you when you return to our website and helping our team to understand which sections of the website you find most interesting and useful.